
诊断学理论与实践 ›› 2026, Vol. 25 ›› Issue (03): 370-376.doi: 10.16150/j.1671-2870.2026.03.014
李斯文, 杜红宇, 陈华琴( )
收稿日期:2024-06-10
修回日期:2024-09-17
接受日期:2024-10-08
出版日期:2026-06-25
发布日期:2026-06-27
通讯作者:
陈华琴 E-mail: acai1969718@126.com作者简介:作者贡献/Authors’ Contributions
李斯文负责实施研究,文章选题,采集数据,分析数据,撰写文章;杜红宇负责实施研究,文章选题,采集数据;陈华琴负责文章选题,分析数据,解释数据,文章修改。
LI Siwen, DU Hongyu, CHEN Huaqin()
Received:2024-06-10
Revised:2024-09-17
Accepted:2024-10-08
Published:2026-06-25
Online:2026-06-27
摘要:
假性甲状旁腺功能减退症(pseudohypoparathyroidism, PHP)是一组由外周靶组织对甲状旁腺激素(parathyroid, PTH)抵抗所致的一组临床综合征,其临床特征主要为低钙血症、高磷血症,和由此引起的手足搐搦、惊厥或癫痫样发作。PHP 1b型患者常无特异性体征,临床表现多样,若不及时完善生化、PTH等检测,易延误诊断。本文报道1例以发作性惊厥为主要表现的PHP患儿,男性,11岁,其血清钙、游离钙均降低,同时PTH明显升高,血磷升高,尿钙、尿磷低,基因检测显示该患儿STX16基因5-7号外显子杂合缺失,GNAS-A/B母源甲基化缺失,检查其父母STX16/GNAS-AS1/GNAS等基因/区域拷贝数变化及甲基化情况,结果均未见异常。该患儿明确诊断为新发STX16基因突变所致常染色体显性遗传型PHP 1b型(autosomal dominant inheritance-PHP 1b, AD-PHP 1b)。予骨化三醇及钙剂口服治疗后,其血钙逐渐恢复正常,未再惊厥发作,目前仍在跟踪随访。目前国内未见报道男性STX16基因新发突变导致AD-PHP 1b型病例。
中图分类号:
李斯文, 杜红宇, 陈华琴. STX16新发突变致假性甲状旁腺功能减退症1b型1例报告[J]. 诊断学理论与实践, 2026, 25(03): 370-376.
LI Siwen, DU Hongyu, CHEN Huaqin. A case report of pseudohypoparathyroidism type 1b caused by noval mutation of STX16[J]. Journal of Diagnostics Concepts & Practice, 2026, 25(03): 370-376.

图1
患儿头颅CT:双侧放射冠、基底节区对称性分布钙化灶


图2
患儿样本(右)与正常对照(左)比对,STX16基因5-7号外显子杂合缺失(RFU值下降)


图3
患儿STX16基因5-7号外显子杂合缺失,GNAS-A/B甲基化缺失


图4
患儿父亲样本(右)与正常对照(左)比对,STX16/GNAS-AS1/GNAS等基因/区域拷贝数未见异常


图5
患儿父亲STX16/GNAS-AS1/GNAS等基因/区域甲基化检测未见异常


图6
患儿母亲样本(右)与正常对照(左)比对,STX16/GNAS-AS1/GNAS等基因/区域拷贝数未见异常


图7
患儿母亲STX16/GNAS-AS1/GNAS等基因/区域甲基化检测未见异常

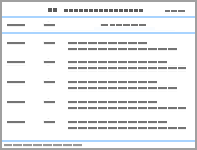
表1
患儿出院后复查相关指标情况
| 时间 | 25-羟维生素D (ng/mL) | 血钙 (mmol/L) | 血磷 (mmol/L) | 甲状旁腺素 (pg/mL) |
|---|---|---|---|---|
| 出院后1个月 | 32.32 | 1.62 | 2.74 | - |
| 出院后1.5个月 | - | 1.80 | 2.71 | 590.8 |
| 出院后2个月 | 37.96 | 1.99 | 2.57 | - |
| 出院后3个月 | 27.09 | 1.95 | 2.4 | - |
| 出院后3.5个月 | 33.28 | 1.89 | 2.28 | - |
| 出院后4个月 | 38.78 | 2.14 | 2.2 | 445.6 |
| 出院后4.5个月 | 35.46 | 2.32 | 2.26 | - |
| 出院后5个月 | 41.46 | 2.35 | 2.13 | 462.7 |
| 出院后7个月 | 48.22 | 2.30 | 1.83 | - |
| 出院后8个月 | 40.54 | 2.36 | 1.55 | 200 |
| 出院后13个月 | 41.18 | 2.15 | 1.72 | 643.9 |
| 出院后14个月 | 39.39 | 2.37 | 1.54 | 373.75 |
| 出院后20个月 | 39.58 | 2.48 | 1.59 | 82.6 |
| 出院后25个月 | 38.13 | 2.41 | 1.54 | 78.2 |
| [1] |
MANTOVANI G, BASTEPE M, MONK D, et al. Diagnosis and management of pseudohypoparathyroidism and related disorders: first international Consensus Statement[J]. Nat Rev Endocrinol, 2018, 14(8):476-500.
doi: 10.1038/s41574-018-0042-0 pmid: 29959430 |
| [2] | 中华医学会骨质疏松和骨矿盐疾病分会,中华医学会内分泌分会代谢性骨病学组. 甲状旁腺功能减退症临床诊疗指南[J]. 中华骨质疏松和骨矿盐疾病杂志, 2018, 11(4):323-337. |
| Chinese Medical Association Osteoporosis and Bone Mine-ral Diseases Branch, Metabolic Osteopathy Group, Endocrinology Branch, Chinese Medical Association. Clinical diagnosis and treatment guidelines for hypoparathyroidism[J]. Chin J Osteoporosis&Bone miner Res, 2018, 11(4):323-337. | |
| [3] | YAVROPOULOU M P, CHRONOPOULOS E, TROVAS G, et al. Hypercalcitoninaemia in pseudohypo-parathyroidism type 1A and type 1B[J]. Endocrinol Diabetes Metab Case Rep, 2019,2019:18-0125. |
| [4] | DIXIT A, CHANDLER K E, LEVER M, et al. Pseudohypoparathyroidism type 1b due to paternal uniparental disomy of chromosome 20q[J]. J Clin Endocrinol Metab, 2013, 98(1):E103-E108. |
| [5] | MANTOVANI G, ELLI F M. Inactivating PTH/PTHrP signaling disorders[J]. Front Horm Res, 2019,51:147-159. |
| [6] |
JÜPPNER H. Molecular definition of pseudohypoparathyroidism variants[J]. J Clin Endocrinol Metab, 2021, 106(6):1541-1552.
doi: 10.1210/clinem/dgab060 pmid: 33529330 |
| [7] | LOID P, PEKKINEN M, REYES M, et al. GNAS, PDE4D, and PRKAR1A mutations and GNAS methylation changes are not a common cause of isolated early-onset severe obesity among finnish children[J]. Front Pediatr, 2020,8:145. |
| [8] |
BRANCATELLA A, MANTOVANI G, ELLI F M, et al. A severe inactivating PTH/PTHrP signaling disorder type 2 in a patient carrying a novel large deletion of the GNAS gene: a case report and review of the literature[J]. Endocrine, 2020, 67(2):466-472.
doi: 10.1007/s12020-020-02195-7 |
| [9] |
GERMAIN-LEE E L. Management of pseudohypoparathyroidism[J]. Curr Opin Pediatr, 2019, 31(4):537-549.
doi: 10.1097/MOP.0000000000000783 URL |
| [10] | 中国研究型医院学会甲状旁腺及骨代谢疾病专业委员会, 中华医学会骨质疏松和骨矿盐疾病分会, 王鸥, 等. 术后甲状旁腺功能减退症管理专家共识[J]. 中华内分泌代谢杂志, 2024, 40(10):817-825. |
| Society of Parathyroid and Metabolic Bone Diseases, Chinese Research Hospital Association; Chinese Society of Osteoporosis and Bone Mineral Research, WANG O, et al. Consensus on the management of postsurgical hypoparathyroidism[J]. Chin J Endocrinol Metab, 2024, 40(10):817-825. | |
| [11] |
GRIGELIONIENE G, NEVALAINEN P I, REYES M, et al. A large inversion involving GNAS Exon A/B and all exons encoding Gsα is associated with autosomal dominant pseudohypoparathyroidism type Ⅰb (PHP1B)[J]. J Bone Miner Res, 2017, 32(4):776-783.
doi: 10.1002/jbmr.3083 URL |
| [12] |
ELLI F M, LINGLART A, GARIN I, et al. The prevalence of GNAS deficiency-related diseases in a large cohort of patients characterized by the EuroPHP network[J]. J Clin Endocrinol Metab, 2016, 101(10):3657-3668.
doi: 10.1210/jc.2015-4310 pmid: 27428667 |
| [13] |
KIUCHI Z, REYES M, HANNA P, et al. Progression of PTH resistance in autosomal dominant pseudohypoparathyroidism type Ⅰb due to maternal STX16 deletions[J]. J Clin Endocrinol Metab, 2022, 107(2):e681-e687.
doi: 10.1210/clinem/dgab660 URL |
| [14] | BRIX B, WERNER R, STAEDT P, et al. Different pattern of epigenetic changes of the GNAS gene locus in patients with pseudohypoparathyroidism type Ⅰc confirm the heterogeneity of underlying pathomechanisms in this subgroup of pseudohypoparathyroidism and the demand for a new classification of GNAS-related disorders[J]. J Clin Endocrinol Metab, 2014, 99(8):E1564-E1570. |
| [15] | 曹莹, 路文盛. 假性甲状旁腺功能减退症Ⅰ型遗传学研究进展[J]. 国际内分泌代谢杂志, 2022, 42(3):184-186,194. |
| CAO Y, LU W S. Progress in genetic research of type Ⅰ pseudohypoparathyroidism[J]. International Journal of Endocrinology and Metabolism, 2022, 42(3):184-186,194. | |
| [16] |
MANTOVANI G, BONDIONI S, LINGLART A, et al. Genetic analysis and evaluation of resistance to thyrotropin and growth hormone-releasing hormone in pseudohypoparathyroidism type Ⅰb[J]. J Clin Endocrinol Metab, 2007, 92(9):3738-3742.
doi: 10.1210/jc.2007-0869 URL |
| [17] | TURAN S, IGNATIUS J, MOILANEN J S, et al. De novo STX16 deletions: an infrequent cause of pseudohypoparathyroidism type Ⅰb that should be excluded in sporadic cases[J]. J Clin Endocrinol Metab, 2012, 97(12):E2314-E2419. |
| [18] |
BASTEPE M, FRÖHLICH L F, HENDY G N, et al. Autosomal dominant pseudohypoparathyroidism type Ⅰb is associated with a heterozygous microdeletion that likely disrupts a putative imprinting control element of GNAS[J]. J Clin Invest, 2003, 112(8):1255-1263.
doi: 10.1172/JCI19159 URL |
| [19] |
LINGLART A, GENSURE R C, OLNEY R C, et al. A novel STX16 deletion in autosomal dominant pseudohypoparathyroidism type Ⅰb redefines the boundaries of a cis-acting imprinting control element of GNAS[J]. Am J Hum Genet, 2005, 76(5):804-814.
doi: 10.1086/429932 URL |
| [20] |
KIUCHI Z, REYES M, JÜPPNER H. Preferential Maternal transmission of STX16-GNAS mutations responsible for autosomal dominant pseudohypoparathyroidism type Ⅰb (PHP1B): another example of transmission ratio distortion[J]. J Bone Miner Res, 2021, 36(4):696-703.
doi: 10.1002/jbmr.4221 URL |
| [21] |
ZAVATTA G, CLARKE B L. Basal ganglia calcification in hypoparathyroidism and pseudohypoparathyroidism: local and systemic metabolic mechanisms[J]. J Endocrinol Invest, 2021, 44(2):245-253.
doi: 10.1007/s40618-020-01355-w |
| [22] |
CHU X, ZHU Y, WANG O, et al. Bone mineral density and its serial changes are associated with PTH levels in pseudohypoparathyroidism type 1B patients[J]. J Bone Miner Res, 2018, 33(4):743-752.
doi: 10.1002/jbmr.3360 pmid: 29240265 |
| [23] | 徐静尊, 尹雪瑶. 1例Ⅰb型假性甲状旁腺功能减退症合并低钾血症的临床和遗传特征分析[J]. 中华内分泌代谢杂志, 2024, 40(6):525-528. |
| XU J Z, YIN X Y. Clinical and genetic analysis of a case of type Ib pseudohypoparathyroidism complicated with hypokalemia[J]. Chin J Endocrinol Metab, 2024, 40(6):525-528. |
| [1] | 王刚, 王玲玉. 遗传性血小板减少症诊断现状及挑战[J]. 诊断学理论与实践, 2026, 25(01): 9-14. |
| [2] | 黄晓燕, 赵小峰, 鲍军明,. 一个先天性双侧无眼症家系的临床调查及分析[J]. 诊断学理论与实践, 2012, 11(04): 394-396. |
| [3] | 胡晓波, 季慧峰, 金大鸣,. 白细胞形态中的Pelger-Huёt异常[J]. 诊断学理论与实践, 2006, 5(04): 370-372. |
| [4] | 汤兵,梅长林,张玲,章建全,孙田美,张树忠. 多囊肾病患者肾脏体积与临床表现关系的研究[J]. 诊断学理论与实践, 2003, 2(04): 43-45. |
| [5] | 梅长林,李林. 常染色体显性遗传性多囊肾病诊治进展[J]. 诊断学理论与实践, 2003, 2(04): 18-20. |
| 阅读次数 | ||||||
|
全文 |
|
|||||
|
摘要 |
|
|||||
